-
69
Ligand geometry-dependent coordination of alkali halide cluster ions by bipyridine isomers in the gas phase (PCCP, 2026)
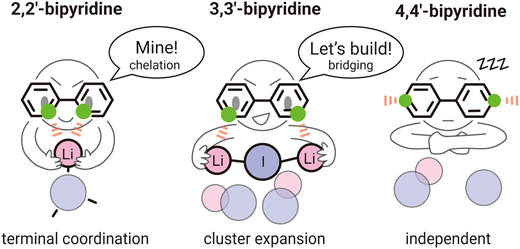
Ligand geometry plays an important role in metal-ion coordination and influences both the structures and growth of ligand-bound alkali halide clusters. Here, we investigate the interactions of LiI cluster cations with three bipyridine isomers using electrospray ionization–ion mobility spectrometry–mass spectrometry (ESI-IMS-MS) combined with density functional theory (DFT) calculations. Ion mobility measurements together with theoretical collision cross section calculations reveal that 2,2′-bipyridine consistently adopts localized bidentate chelation, whereas 3,3′-bipyridine preferentially bridges multiple Li+ centers. These distinct coordination modes give rise to fundamentally different cluster growth behaviors: chelating coordination limits cluster growth, while bridging coordination stabilizes progressively larger LiI frameworks and can even alter the preferred structures of the ionic cores. In contrast, 4,4′-bipyridine exhibits only weak coordination because of its unfavorable donor geometry. Comparison with our previous study on flexible diamine ligands further reveals that ligand flexibility and donor geometry represent two complementary factors governing ligand-assisted ionic cluster assembly. These results establish a direct relationship between ligand geometry, coordination mode, and ionic cluster architecture, providing molecular-level insight into ligand-directed alkali halide cluster assembly.
Yunseop Choi
Jongcheol Seo*
-
68
Facile synthesis of large-Stokes-shift giant CdHgSe/CdHgSeS near-infrared quantum dots via chlorine-mediated Ostwald ripening (Mater. Today Adv., 2026)
Giant core/shell quantum dots (GQDs) offer advantages for optoelectronics, including suppressed blinking and reduced Auger recombination, yet thick-shell growth often requires laborious successive ion layer adsorption and reaction (SILAR) cycles. Here we demonstrate a chlorine-mediated controlled Ostwald ripening strategy that transforms NIR-emitting CdHgSe/CdS core/shell seed QDs into giant CdHgSe/CdHgSeS core/gradient-shell QDs in a single post-synthetic step. Seed QDs are prepared by growing a thin (∼1.6-monolayer) CdS shell on CdHgSe alloy cores. Injection of CHCl3 at 240 °C triggers rapid ripening, in which most seeds dissolve while a minor population grows into GQDs. During ripening, the band-edge absorption feature is strongly attenuated and absorption below ∼610 nm increases markedly, whereas NIR emission remains centered near ∼900 nm, yielding a large effective Stokes shift that can mitigate reabsorption in luminescent solar concentrators. HAADF-STEM/EDS and XRD support a compositionally graded shell, while mass spectrometry and numerical simulations are consistent with chlorine-modified surface chemistry and altered monomer solubility/surface tension that accelerate ripening. As a proof of concept, embedding the GQDs in PMMA affords a transparent luminescent solar concentrator prototype with retained NIR emission and favorable spectral overlap with silicon photovoltaics.
Gyudong Lee
Sanghwang Park
Jongcheol Seo
Wonseok Lee
Opeyemi H. Arogundade
Andrew M. Smith
Jongmin Choi
Hongsoo Choi
Sung Jun Lim*
-
67
Extraction and usage of free electrons from alkali metals at room temperature (Comm. Mater., 2026)
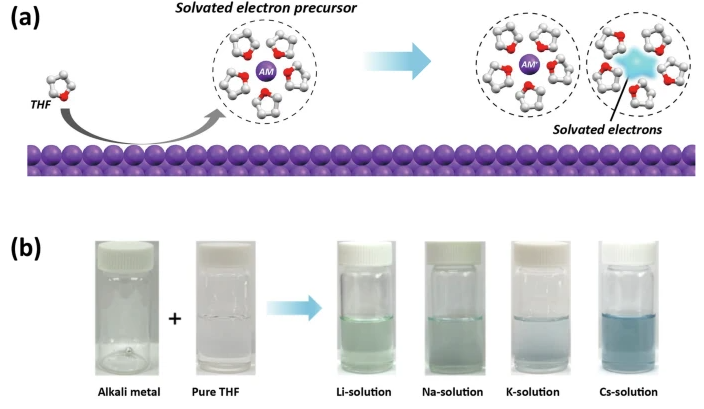
Extracting free electrons from metals is challenging but important for fundamental metal chemistry and future energy applications. The main challenges are electron extraction and stabilization for storage and use. While solvated electrons in liquid ammonia offer a promising approach, their preparation requires harsh conditions below −33.6 °C. Here, we report spontaneous alkali metal (AM) dissolution in tetrahydrofuran (THF) at room temperature, generating stable free-electron solutions. Li, Na, K, and Cs readily dissolve in THF—a widely used organic solvent—producing AM-solutions containing solvated electrons. Mass spectrometry reveals formation of [AM(THF)n]+ complexes (n = 4-10), while X-ray photoelectron spectroscopy confirms the presence of solvated electrons and oxidized metal cations. The favorable reduction potential of THF enables spontaneous AM oxidation, with THF simultaneously stabilizing both extracted electrons and metal cations. We successfully employed AM-solutions for C60 fullerene doping, producing superconducting phases and demonstrating practical utility for materials synthesis. This room-temperature approach overcomes the cryogenic limitations of ammonia-based systems, offering a straightforward method for generating and utilizing solvated electrons under ambient conditions.
Changmin Choi
Sanghwang Park
Yunseop Choi
Jin Young Koo
Taekyung Yoon
Jongcheol Seo*
Hee Cheul Choi*
-
66
Ligand Counting and Size Determination of Quantum Dots Using Electronegative Matrix-Assisted Laser Desorption Ionization Mass Spectrometry (Anal. Chem., 2026)
Surface-capping ligands on quantum dots (QDs) modify the physical and chemical properties of QDs to enable diverse applications. Accurate qualitative and quantitative analyses of these ligands are therefore essential for the advancement of QD research; however, conventional methods (e.g., UV–vis, TEM) seldom provide direct stoichiometric insight. MS can offer a solution, but traditional MALDI induces an excessive ligand loss. Here, we employed a highly electronegative, fluorinated porphyrin (TPFP) as a soft MALDI matrix for CdSe QDs to preserve the ligands and yield high ion signals at reduced laser power. Comparative studies with electron-transfer matrixes (DCTB, 9-nitroanthracene, dithranol, and pentacene) indicate that the highly electronegative TPFP extracts electrons from photoexcited QDs, lowering ionization thresholds and preventing ligand detachment. Using TPFP-MALDI-MS, we determined QD core sizes and counted the number of surface ligands in carboxylate- and thiolate-capped QDs, and for thiolate-capped QDs in particular, we further evaluated ligand numbers and ligand densities with various core sizes. This approach provides a reliable strategy to determine QD core sizes, to count ligand numbers, and to estimate the surface coverage with improved resolution. These findings underscore the importance of electron affinity of a matrix for soft ionization and expand MS-based tools for probing QD core and surface chemistry, paving the way for more reliable characterization of nanocrystalline systems.
Sanghwang Park
Dongyeon Ro
Gyudong Lee
Jinhyuk Ma
Taiha Joo
Chang Hyuck Choi
Seung Koo Shin*
Sung Jun Lim*
Jongcheol Seo*
-
65
Solvation of lithium iodide cluster cations via bridging diamine coordination (JCP, 2025)
Diamines are well-known bidentate ligands that coordinate with metal cations; however, their solvation behavior with ionic clusters remains largely unexplored. In this study, we investigated the structures of lithium iodide ionic clusters solvated by ethylenediamine and N,N,N′,N′-tetramethylethylenediamine using electrospray ionization(ESI)–ion mobility spectrometry–mass spectrometry. Interestingly, diamines were found to stabilize ionic clusters by bridging two metal cations, even though such structures are not the most thermodynamically stable in the gas phase. We propose that these bridging structures were likely formed and kinetically trapped in the solution phase or in the highly concentrated droplets generated during the ESI process. Furthermore, we analyzed the effects of alkali metal and halide identity on solvation behavior.
Yunseop Choi
Junyoung Ahn
Minsu Kim
Jongcheol Seo*
-
64
Inducing a synergistic anti-obesity effect by increasing the bioavailability of the flavonoid rutin with a L. plantarum strain (Food Chemistry, 2025)
Rutin, a sugar-bonded flavonoid, exhibits anti-obesity potential but has limited bioavailability unless hydrolyzed to isoquercetin or further to quercetin. This study evaluated Lactiplantibacillus plantarum HAC03 (HAC03), isolated from Korean white kimchi, as an effective facilitator of rutin bioavailability. HAC03 rapidly converted rutin into isoquercetin and, at a slower rate, to quercetin both in vitro and in vivo. Unlike HAC03-only or rutin-only groups, co-administration of HAC03 and rutin synergistically reduced body weight, adipocyte size, and obesity biomarkers by suppressing fat synthesis and enhancing fatty acid β-oxidation in diet-induced obese mice. HAC03, together with rutin, colonized the ileum and enriched polyphenol-metabolizing gut microbes. At 30 min post-administration, isoquercetin—more bioavailable and effective than its precursor, rutin—peaked (∼9 μM) in mouse portal vein blood, while remaining minimal or undetectable in controls. This pairing of functional flavonoid and tailored probiotic synergistically combats obesity via ileal isoquercetin conversion, offering a promising single-flavonoid, single-probiotic obesity-management strategy.
Youn-Goo Kang
Seongjae Jang
Bobae Kim
Jongcheol Seo
Ah-Ram Kim*
-
63
Use of Carbodicarbene as a Single Organic Initiator in the Conjugate-Addition Polymerization of (Meth)acrylic Monomers (J. Polym. Sci., 2025)
To enable efficient anionic polymerization without metal catalysts and co-modulators, conjugate-addition polymerization of (meth)acrylic monomers was conducted using carbodicarbene (CDC) coordinated with N-heterocyclic carbene and cyclic (alkyl)(amino)carbene. As the sole organic initiator, CDC showed high activity and polymerization efficiency when used with different (meth)acrylic monomers, including methacrylates with functional groups. The role of CDC as an initiator was supported by nanoelectrospray ionization mass spectrometry.
Kunou Kim
Minseop Kim
Yoonjin Kim
Jongcheol Seo
Eunsung Lee*
Kyung-sun Son*
-
62
Structures and energetics of protonated bipyridine–cucurbituril complex isomers in the gas phase (PCCP, 2025)
We investigated the gas-phase isomerism and stability of host–guest complexes formed between cucurbiturils (CB[6] and CB[7]) and three n,n′-bipyridine regioisomers (n = 2, 3, and 4), focusing on how molecular geometry and charge distribution influence complex formation. Ion mobility spectrometry-mass spectrometry and collision-induced dissociation experiments, supported by density functional theory (DFT) calculations, reveal distinct inclusion and exclusion complex isomers. Singly-protonated bipyridines tend to form exclusion complexes with CB[6], while doubly-protonated forms enable stable inclusion through enhanced charge–portal interactions. CB[7], with its larger and more flexible cavity, consistently supports inclusion for all bipyridine isomers, regardless of charge state. These findings emphasize the importance of charge localization, host flexibility, and phase-specific effects in supramolecular assembly, which may further offer valuable insights for designing bipyridine- or bipyridinium-based materials.
Doui Kim
Jongcheol Seo*
-
61
Spin‐State and Clustering Effects in Fe‐Complex Negolytes for Near‐Neutral Aqueous Redox Flow Batteries (Angew. Chem., 2025)
Cost-effective redox-active materials are essential for advancing redox flow batteries (RFBs). Iron, with its abundance and suitability as a redox couple, is a promising candidate; however, achieving stable and fast redox reactions in aqueous RFBs remains a challenge. This study presents an Fe-based negolyte stabilized by a hexadentate ligand, where Fe–ligand bonds are enhanced through intermolecular interactions. The sulfonate-substituted Fe complex exhibits a formal potential of −0.44 V versus Ag/AgCl and an exceptionally high rate constant of 0.69 cm s−1. Near-neutral RFBs incorporating 0.5 M Fe complex show excellent cycling stability, with no discernible capacity fading over 300 cycles. This performance is attributed to intermolecular hydrogen bonds that reinforce Fe–ligand coordination and promote the formation of stable trimeric clusters. Operando electrochemical Raman spectroscopy and density functional theory reveal that π-backdonation from Fe(II) to the imino-phenolate moiety further stabilizes the complex after reduction. In contrast, the hydroxyl-substituted complex exhibits inferior stability due to weaker hydrogen bonding and less pronounced π-backdonation. These findings underscore the importance of ligand design and intermolecular interactions in developing cost-effective, high-performance redox-active materials for aqueous RFBs.
Donghwi Ko
Seongyeon Kwon
Jantakan Nedsaengtip
Yohan Kim
Yunseop Choi
Dongwook Kim
Xingyi Lyu
Ruchi Dixit
Yugang Zhang
Tao Li
Jongcheol Seo
Mu‐Hyun Baik*
Hye Ryung Byon*
-
60
Expanded ribosomal synthesis of non-standard cyclic backbones in vitro (Nat. Commun., 2025)
The ribosome polymerizes L-α-amino acids into polypeptides, catalyzing peptide bond formation through aminolysis. This process is facilitated by entropy trapping within its peptidyl transferase center (PTC). In this research, we harness this capability to synthesize polymers containing cyclic motifs in the backbone. We design 26 non-canonical monomers (ncMs) with two distinct substrates: dicarboxylic esters and hydrazinoesters, each containing bifunctional moieties that undergo ring-closing reactions through multiple aminolysis reactions. Using a cell-free system that enables the consecutive incorporation of these ncMs into a growing peptide, we discover that the ribosome can produce 5- and 6-membered cyclic backbones, which have never been reported. We also demonstrate that the formation of such cyclic backbones within the ribosome is tunable by altering the substituents of dicarboxylic esters. This discovery expands the range of non-standard backbones that can be synthesized by the ribosome and motivates future research towards expanding ribosome-mediated chemistries for biopolymer synthesis.
Kanghun Lee
Hyeongwoo Park
Ravi Kumar Devarapalli
Dahye Im
Jongcheol Seo
Joongoo Lee*

 68Facile synthesis of large-Stokes-shift giant CdHgSe/CdHgSeS near-infrared quantum dots via chlorine-mediated Ostwald ripening (Mater. Today Adv., 2026)Giant core/shell quantum dots (GQDs) offer advantages for optoelectronics, including suppressed blinking and reduced Auger recombination, yet thick-shell growth often requires laborious successive ion layer adsorption and reaction (SILAR) cycles. Here we demonstrate a chlorine-mediated controlled Ostwald ripening strategy that transforms NIR-emitting CdHgSe/CdS core/shell seed QDs into giant CdHgSe/CdHgSeS core/gradient-shell QDs in a single post-synthetic step. Seed QDs are prepared by growing a thin (∼1.6-monolayer) CdS shell on CdHgSe alloy cores. Injection of CHCl3 at 240 °C triggers rapid ripening, in which most seeds dissolve while a minor population grows into GQDs. During ripening, the band-edge absorption feature is strongly attenuated and absorption below ∼610 nm increases markedly, whereas NIR emission remains centered near ∼900 nm, yielding a large effective Stokes shift that can mitigate reabsorption in luminescent solar concentrators. HAADF-STEM/EDS and XRD support a compositionally graded shell, while mass spectrometry and numerical simulations are consistent with chlorine-modified surface chemistry and altered monomer solubility/surface tension that accelerate ripening. As a proof of concept, embedding the GQDs in PMMA affords a transparent luminescent solar concentrator prototype with retained NIR emission and favorable spectral overlap with silicon photovoltaics.Gyudong Lee Sanghwang Park Jongcheol Seo Wonseok Lee Opeyemi H. Arogundade Andrew M. Smith Jongmin Choi Hongsoo Choi Sung Jun Lim*